CorrFlow is proposed, a correlation-guided flow matching framework for histology-to-ST prediction that explicitly models gene-gene dependencies through two complementary mechanisms and introduces an annealed masked flow matching strategy, encouraging the model to infer masked genes conditioned on the remaining genes and promoting joint conditional modeling beyond per-gene marginal estimation.
Abstract
Spatial transcriptomics (ST) provides spatially resolved gene expression profiling but remains expensive, motivating the prediction of ST from histology images. Generative models have emerged as a mainstream paradigm for ST prediction due to their ability to model the conditional distribution of gene expression and capture its inherent stochasticity. However, these methods typically treat genes as independent prediction targets and overlook the intrinsic gene-gene interactions in biological systems, which limits their ability to preserve biologically meaningful co-expression patterns. We argue that gene-gene interactions, which reflect shared pathways and regulatory mechanisms, are essential for generating numerically accurate and biologically coherent ST profiles. In this paper, we propose CorrFlow, a correlation-guided flow matching framework for histology-to-ST prediction that explicitly models gene-gene dependencies through two complementary mechanisms. First, we introduce an annealed masked flow matching strategy, where subsets of genes are progressively masked following a timestep-dependent annealing schedule, encouraging the model to infer masked genes conditioned on the remaining genes and promoting joint conditional modeling beyond per-gene marginal estimation. Second, we devise a gene graph-regularized optimization scheme that integrates prior knowledge from the STRING database and data-driven co-expression estimated by WGCNA to construct a gene affinity graph, which enforces both local consistency and global smoothness in the predicted expression. Extensive experiments across 12 datasets show that CorrFlow achieves the best average PCC and HPCC among evaluated methods, leading to more biologically coherent ST predictions.

Spatial transcriptomics enables high-throughput gene expression profiling while preserving spatial information, offering valuable insights into tissue architecture and cellular organization. Methods that achieve single-cell or subcellular spatial resolution typically rely on predefined gene panels, which limits genome-...
A. Abir, Muhtasim Noor Alif, Wei Zhang· bioRxiv· 0 citations
Imaging-based spatial transcriptomics enables molecule-resolved profiling of gene expression and tissue organization in situ. However, segmentation errors, transcript spillover and three-dimensional cell overlap can introduce misassigned transcripts into cell-level expression profiles, compromising biological interpret...
Spatial transcriptomics enables gene expression profiling within intact tissues while preserving spatial context, providing unprecedented insights into cellular organization and function. However, mRNA diffusion during tissue processing can cause transcripts originating from adjacent cells to be captured at a given spo...
Ji-Xin Liu, Shu-Li Sun, Yang Xu et al.· bioRxiv· 0 citations
This work introduces SpaFactor, a lightweight and efficient low-rank morphology-program-gene factorization framework that demonstrates that lightweight joint modeling of tissue context and gene programs can improve both predictive accuracy and biological fidelity.
Shi-Ting Ruan, Xi-Tong Ling, Qi-Ming He et al.· arXiv.org· 0 citations
Spatial transcriptomics (ST) enhances single-cell RNA sequencing by revealing transcript distribution, offering critical insights into heterogeneous diseases such as breast cancer. However, the high cost and lengthy processes of generating high-quality ST data limit clinical application. Recent deep learning methods pr...
CLEAR-ST provides an interpretable and practical approach for diffusion-aware correction of capture-based spatial transcriptomics data, and improved spatial domain recovery, increased gene-level spatial autocorrelation, and enhanced the biological specificity of downstream analyses such as marker gene discovery, pathwa...
K. Ma, Yuan-Hua Huang, J. Ho· bioRxiv· 0 citations
Related blog posts
MIT News · Artificial Intelligence· news.mit.eduOct 28, 2026
Professor Paris Smaragdis develops computers with superhuman hearing for applications like speech recognition, music understanding, and environmental monitoring.
Exploring how generative AI could make machine vision more accessible to businesses. The post GenEye in a Box: Making Machine Vision Something You Can Just Ask For appeared first on GPT-Lab.
Microsoft Research Blog· microsoft.comAug 11, 2026
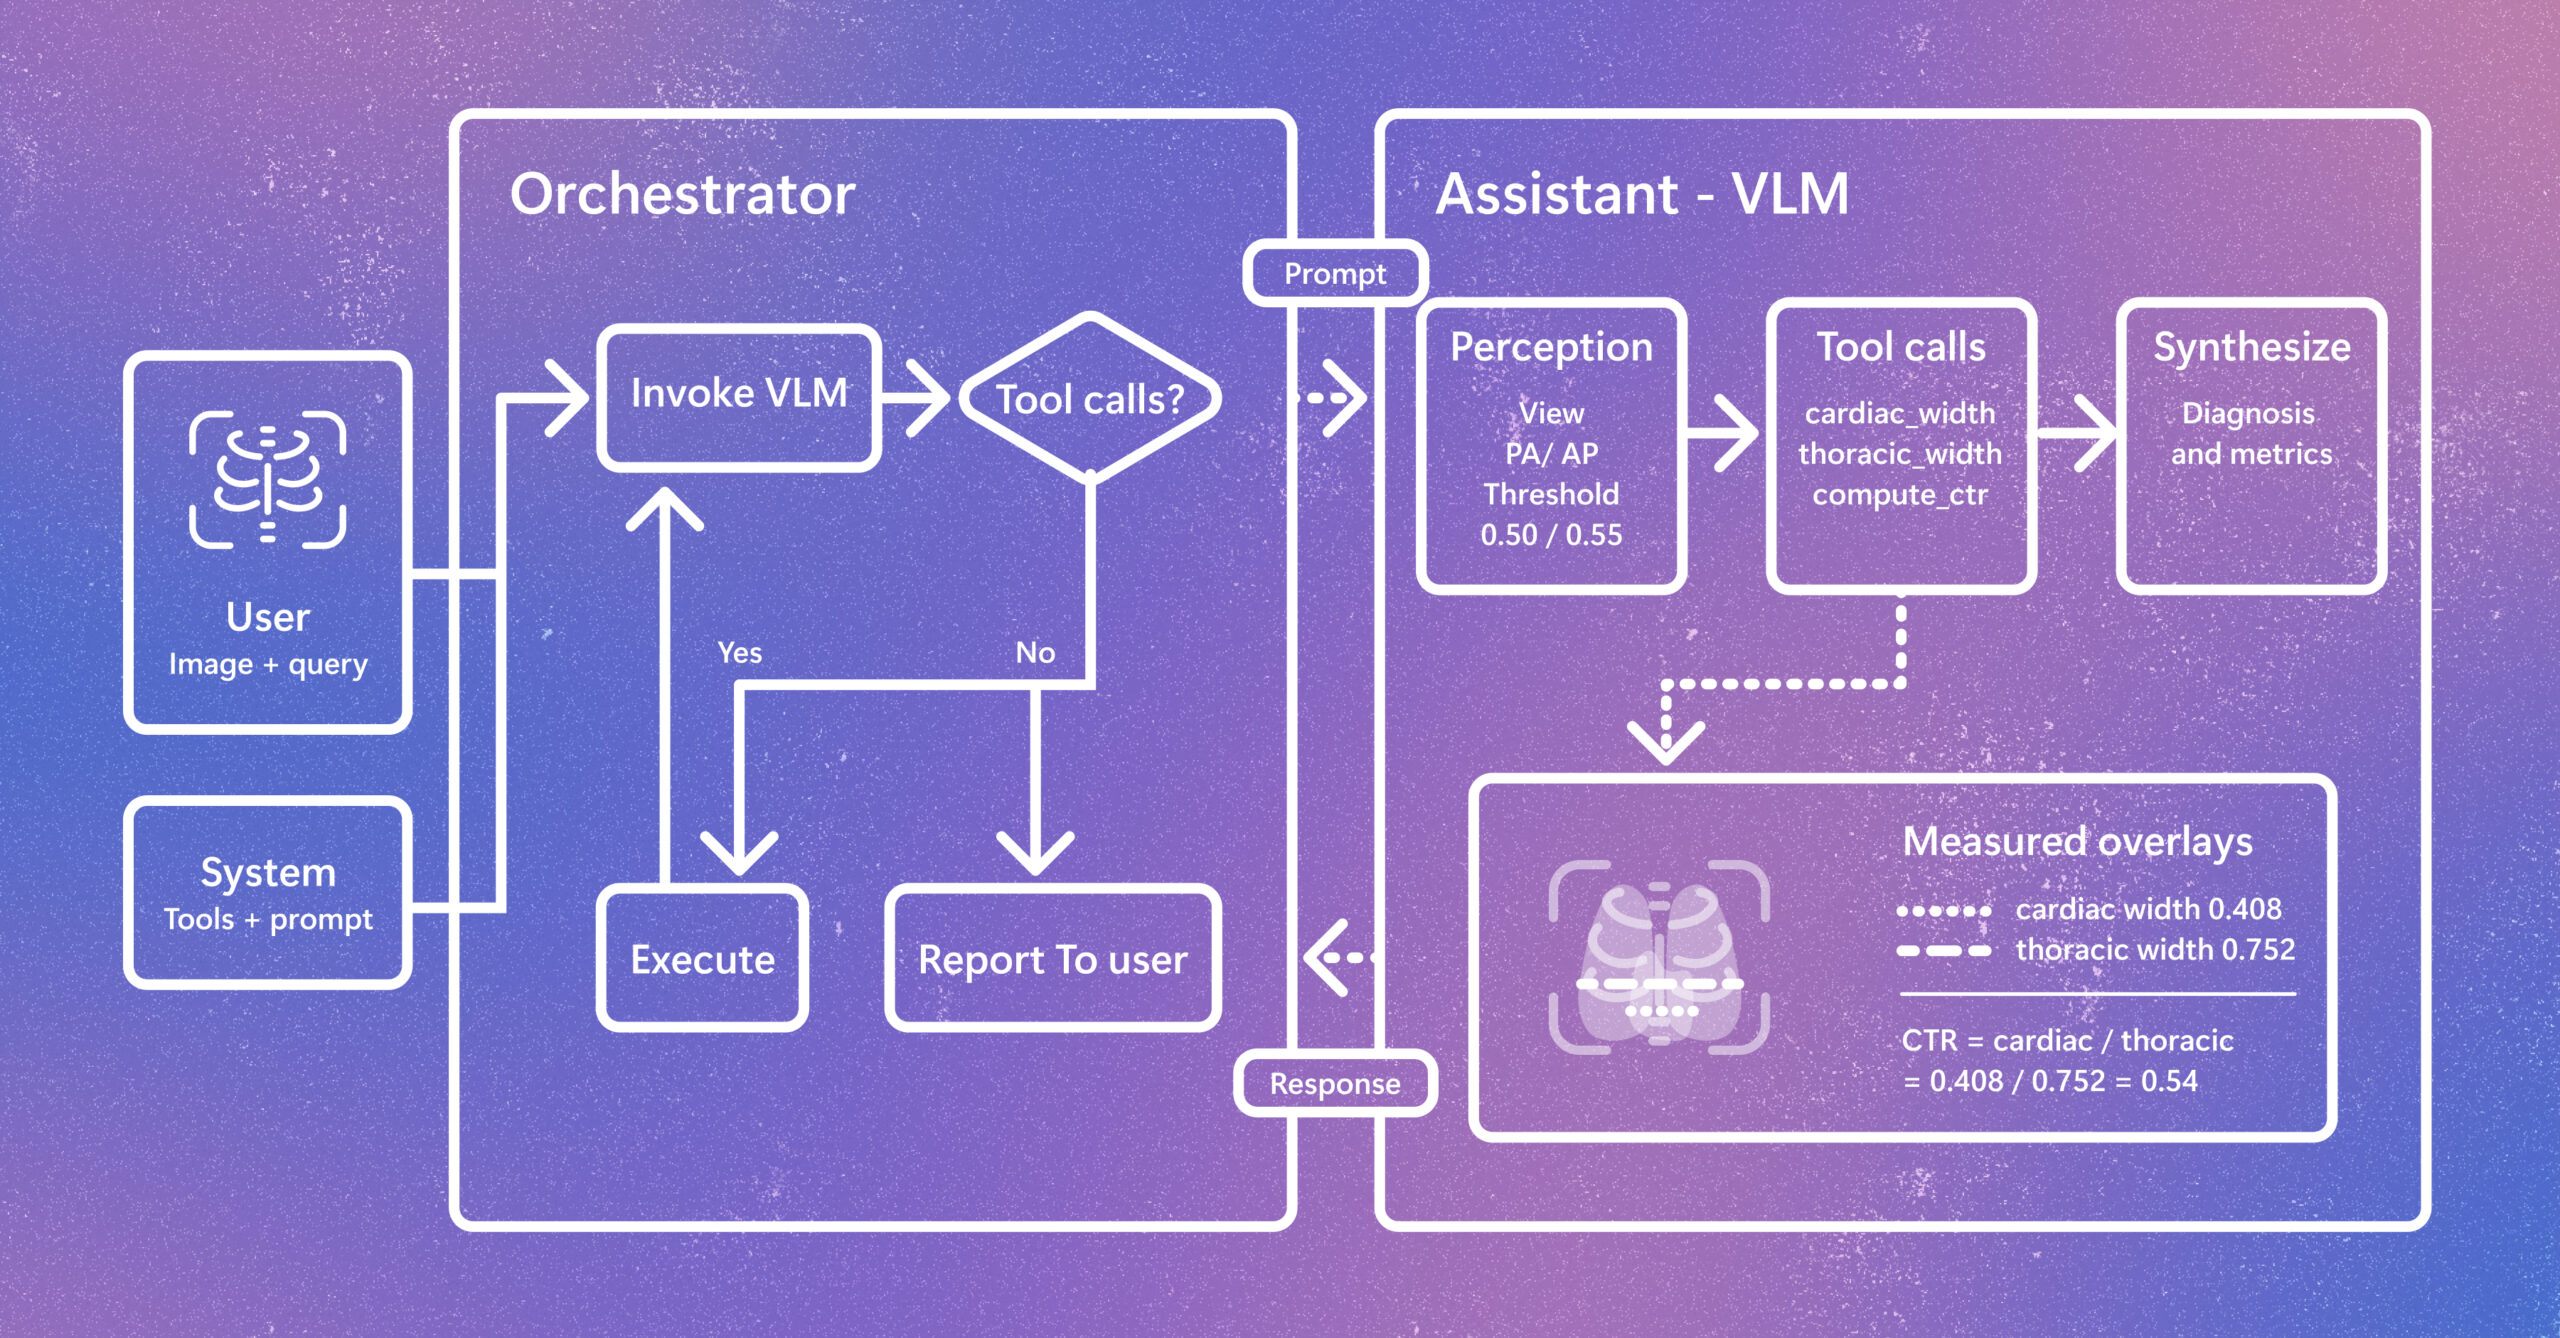
Radiology AI is evolving beyond report generation. CARE-X explores a unified approach that combines flexible reasoning, calibrated predictions, and measurement-based tools for chest X-ray interpretation. The post Introducing CARE-X: Towards Clinically Useful Radiology VLMs with Auxiliary Supervision, Reward-Aligned Learning, and Tool-Augmented Measurement appeared first on Microsoft Research.