It is demonstrated that hotspot-guided design strategy, previously applied to proteins and d-peptides, can be effectively extended to l-peptide scaffolds, enabling the development of functional HA inhibitors informed by key residues at antigen–antibody interfaces.
Abstract
Here we report the applicability of a hotspot-centric approach for de novo computational design of l-peptides based on crystal structures of antigen–antibody complexes. Broadly neutralizing antibodies (bnAbs) FI6v3 and CR9114, which target influenza A virus hemagglutinin (HA), guided the peptide design. Disembodied hotspot residues from these bnAbs were selected and anchored onto known peptide scaffolds (≤35 amino acids) from the Protein Data Bank (PDB). The top-scoring designs were subsequently synthesized, folded, tested in vitro, and structurally characterized in complex with HA. The designed peptides demonstrate structural and functional mimicry by recapitulating the binding mode of the bnAbs on HA and act as inhibitors of low-pH-dependent conformational transitions in HA that facilitate membrane fusion. These findings demonstrate that hotspot-guided design strategy, previously applied to proteins and d-peptides, can be effectively extended to l-peptide scaffolds, enabling the development of functional HA inhibitors informed by key residues at antigen–antibody interfaces.
It is discovered that peptide inhibitors with the wildtype sequence of the heptad repeat 2 (HR2) domain of the SARS-CoV-2 spike protein are efficiently cleaved by Transmembrane Protease, Serine 2, a key protease involved in the SARS-CoV-2 virus-cell fusion pathway.
Kailu Yang, Chu-Chu Wang, Francesco Topi et al.· bioRxiv· 0 citations
Detailed molecular dynamics simulations combined with free energy perturbation calculations were used to systematically investigate the effects of point mutations on both the fusion peptide and the antibody, providing a quantitative and structural framework for understanding fusion peptide recognition.
Mei Feng, Nan Wang, Yi Song et al.· Journal of Physical Chemistr...· 0 citations
SARS-CoV-2 papain-like protease (PLpro) remains an attractive antiviral target because of its dual role in viral polyprotein processing and host immune modulation. In this study, we employed an exploratory structure-based computational workflow to identify antimicrobial peptide scaffolds with predicted PLpro-binding po...
Erenilson dos Santos Moreira, S. Siqueira, F. Barbosa et al.· Scientific Reports· 0 citations
The objective of the present study was to identify cyclic peptides CycPept_3468 and CycPept_4348, which showed excellent inhibitory activity against the MPXV A42R profilin-like protein, indicating their potential efficacy as antiviral drugs.
Ahmad Firoz, Hani S. H. Mohammed Ali, I. Ullah· Current Drug Targets· 0 citations
The SARS-CoV-2 envelope protein (protein E) is a small but multifunctional structural protein that plays a critical role in viral assembly, budding, and pathogenesis, making it an attractive target for therapeutic intervention. This study investigated the interaction between the SARS-CoV-2 envelope protein (wild type a...
Afifah Husni Riani, Surya Rosa Putra, Devy Maulidya Cahyani et al.· EPJ Web of Conferences· 0 citations
The Zika virus NS2B/NS3 protease is an important antiviral target, and macrocyclic peptide inhibitors represent promising scaffolds because they combine multibasic recognition motifs with conformational restriction. Here, we investigated two structurally related cyclic peptide inhibitors, referred to as CP1 and CP2, wh...
Camilla Vitoria Silva Marinho, M. D. D. de Oliveira, Anderson H. Lima· Journal of Molecular Modelin...· 0 citations
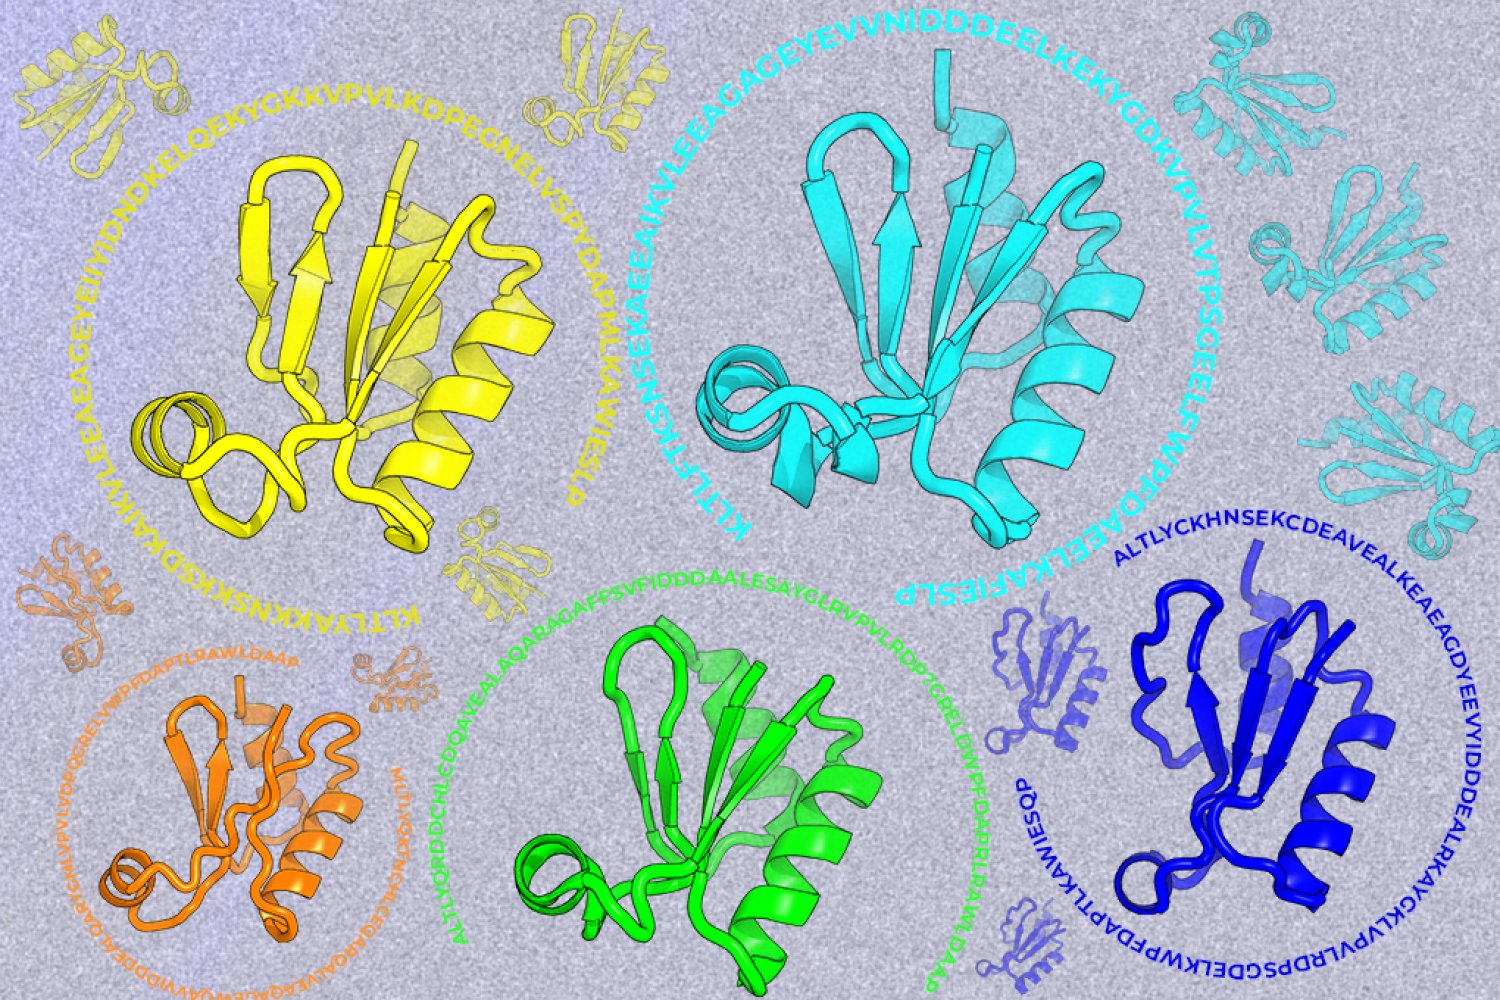
A new machine-learning framework aims to improve the success rate of computational protein design while moving away from results that reproduce sequences found in nature.